Epoxide Reactions: Acid-Assisted Ring Opening (H₃O⁺, ROH/H⁺, HX)
Brønsted acids such as H₃O⁺ protonate epoxides, polarising the C–O bonds so the more substituted carbon carries greater positive character. Weak or neutral nucleophiles (H₂O, ROH, RCO₂H) attack from the backside; attack at the more substituted carbon is often favored by developing positive charge. Regioselectivity depends on the substrate, nucleophile, and conditions. Opening is anti, with inversion when the attacked carbon is stereogenic. Stronger nucleophiles such as halides in HX follow the same anti pathway; they usually favour the less hindered carbon unless a tertiary or benzylic site offers superior stabilisation. The figures below use 2,2-dimethyloxirane to show attack at its tertiary epoxide carbon. The products in these examples are saturated, acyclic structures without a constrained relative geometry, so cis/trans terminology does not apply. They are also achiral.
Quick Summary
- Reagents/conditions: H₃O⁺ (dilute mineral acids), ROH/H⁺ (alcoholysis), or HX (HCl/HBr/HI; often anhydrous).
- Regioselectivity: With H₂O/ROH/RCO₂H, attack often favors the carbon that better stabilizes developing positive charge, especially a tertiary or benzylic site; this is a tendency, not a universal substitution-count rule. Halides in HX default to the less hindered carbon unless a tertiary/benzylic site stabilizes positive charge.
- Stereochemistry: Backside attack on the protonated epoxide gives anti opening and inversion at a stereogenic attacked carbon. Appropriate cyclic epoxides give trans products; acyclic products need not have a cis/trans designation.
- Products: Vicinal diols (H₂O), alkoxy alcohols (ROH), halohydrins (HX), or acyloxy alcohols (RCO₂H); anti opening gives trans substituents when the product ring supports that relationship.
- No rearrangements: The opening is concerted (SN1/SN2 hybrid); discrete carbocations and 1,2-shifts are rare under standard acidic conditions.
Mechanism — Acid-Assisted Epoxide Opening (4 Frames; arrows A–F)
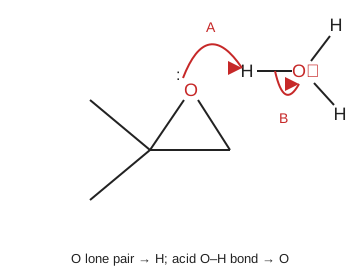
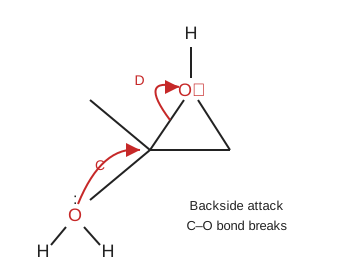
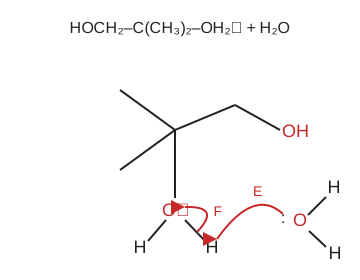
Mechanistic Checklist (Exam Focus)
- Protonate first; the oxonium directs the site of attack.
- With neutral nucleophiles, greater stabilization of developing positive charge often directs attack, particularly at tertiary/benzylic sites; inspect the specific substrate and conditions.
- HX: halide attack is still backside/anti. Less substituted carbons react fastest unless a tertiary/benzylic position stabilises positive charge.
- Anti opening inverts a stereogenic attacked carbon. Use trans for suitable cyclic products, not for every acyclic product.
- Ring strain drives the opening; larger cyclic ethers do not react nearly as readily under comparable conditions.
Worked Examples
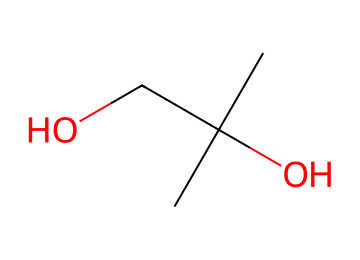
2,2-Dimethyloxirane + H₃O⁺ → 2-methylpropane-1,2-diol
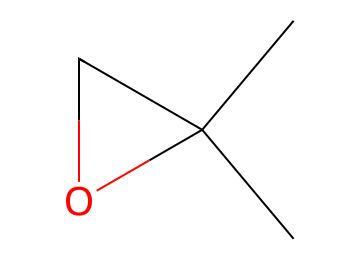
Protonation by H₃O⁺ makes the tertiary carbon most electrophilic, so H₂O adds there. Deprotonation gives HOCH₂C(OH)(CH₃)₂. The two methyl groups are attached to the tertiary alcohol carbon. With unlabeled oxygen atoms, the diol alone does not reveal which oxygen came from water.
2,2-Dimethyloxirane + HBr → 2-bromo-2-methylpropan-1-ol
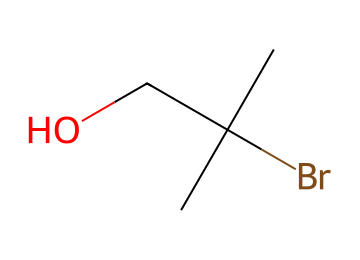
Even though halides often choose the less substituted carbon, the tertiary site here wins because the protonated ring can stabilise the developing cation. The product is HOCH₂C(Br)(CH₃)₂, with Br at the tertiary carbon and OH at the adjacent CH₂. It is achiral.
2,2-Dimethyloxirane + MeOH/H⁺ → 2-methoxy-2-methylpropan-1-ol
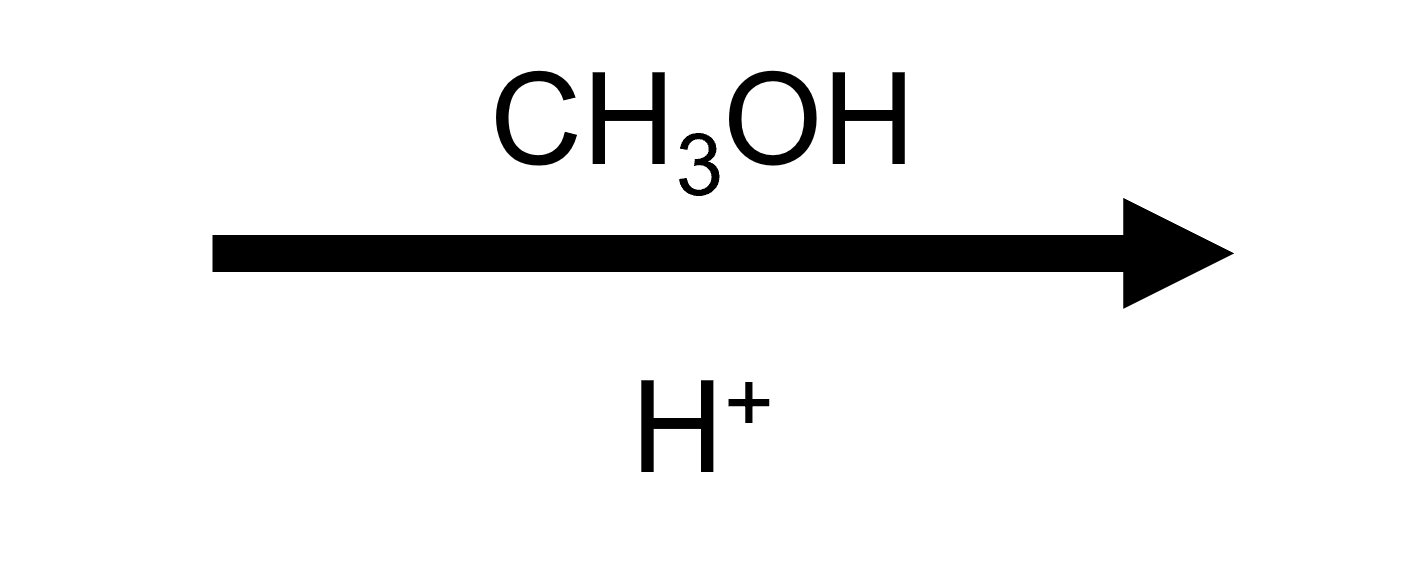
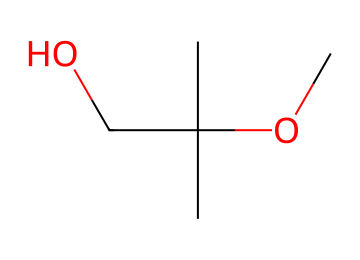
Methanol attacks just like water but leaves behind a protonated OR group. Deprotonation gives HOCH₂C(OMe)(CH₃)₂. The methoxy group is on the carbon bearing both methyl groups; the product is achiral.
2,2-Dimethyloxirane + EtOH/H⁺ → 2-ethoxy-2-methylpropan-1-ol
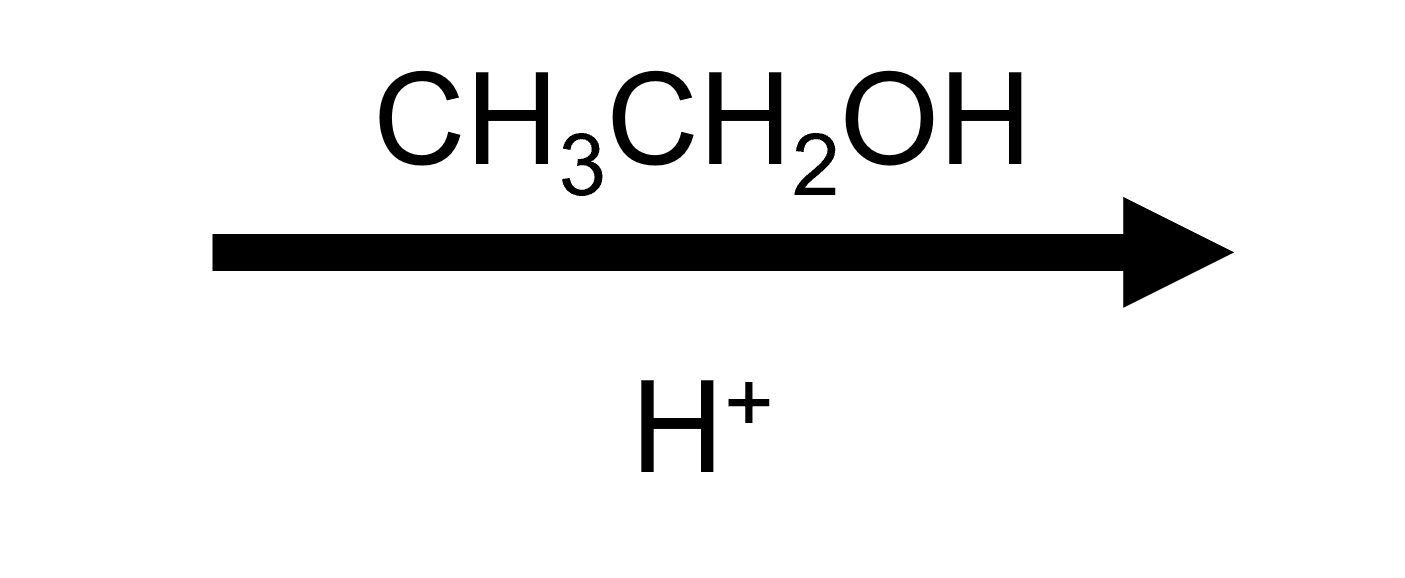
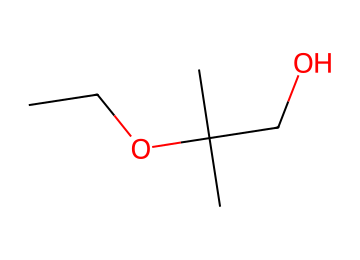
Ethanol gives HOCH₂C(OEt)(CH₃)₂ after deprotonation, again an achiral product. Compare the two to emphasise how changing ROH swaps the alkoxy group in the product.
Scope & Limitations
- Favourable substrates: Aryl, benzylic, allylic, and simple alkyl epoxides. Tertiary centres strongly direct attack to themselves under acidic conditions.
- Regiochemical rules: Neutral nucleophiles often favor the site with greater developing positive-charge stabilization. Halides favor less substituted carbons in simple primary/secondary cases; tertiary/benzylic sites can reverse that preference.
- Cyclic epoxides: Opening gives trans-1,2-disubstituted products; chair conformations dictate axial/equatorial placement.
- Acid-sensitive groups: Protect or avoid strongly acid-labile functionality (acetals, some protecting groups) when using Brønsted acids.
- HF variants: Fluorohydrin synthesis often requires specialized HF complexes (HF·pyridine); adjust conditions accordingly.
- Amines: Protonated amines are poor nucleophiles under these conditions—use basic openings for azide/amine capture instead.
Practical Tips
- Use dilute strong acids (0.1–1 M H₂SO₄, HClO₄, TsOH) with ample H₂O/ROH to minimize rearrangements or polymerization.
- For halohydrins, employ anhydrous HX and keep the medium as dry as practical so the halide remains a strong nucleophile.
- In cyclic systems, draw chair conformations to show anti attack from the less hindered face.
- Quench strongly acidic mixtures cautiously and neutralize residual acid before workup. Capture halogen waste appropriately.
Exam-Style Summary
Epoxide activation by protonation polarizes the ring, pushing positive character onto the more substituted carbon. Backside nucleophilic attack opens the ring with inversion to give overall anti products. H₂O/ROH/RCO₂H often favor the site that better stabilizes developing positive charge; halides usually attack the less hindered carbon unless a tertiary/benzylic site is present. Products are vicinal diols, alkoxy alcohols, or halohydrins. Anti opening gives trans substituents in suitable cyclic products; the achiral examples above have no cis/trans label.
Interactive Toolbox
- Mechanism Solver — Explore supported reactions and compare the result with the protonation, attack, and deprotonation steps above.
- Reaction Solver — Explore available epoxide-opening reagents; check the product against the substrate-specific regioselectivity above.
- IUPAC Namer — Learn the naming ins and outs of epoxide starting materials and their ring-opening products.
Related Reading
- Compare acid-assisted opening with base-promoted epoxide ring opening, where nucleophilic attack favors the less hindered carbon.