Alkene Reactions: Anti Dihydroxylation using RCO₃H and H₃O⁺
Peracids such as mCPBA transform alkenes into epoxides via the Prilezhaev epoxidation, and those rings can then be opened under acidic conditions to furnish vicinal diols by anti addition. The peracid donates an oxygen atom in a concerted transfer, and the resulting epoxide is protonated by hydronium. Water then attacks from the backside (anti to the epoxide oxygen), with the preferred site depending on the epoxide substitution pattern. Both original alkene carbons acquire OH, so the unlabeled diol is not classified as a Markovnikov regioisomer. A final deprotonation gives the neutral anti diol. The standard aqueous hydrolysis proceeds by backside ring opening, giving predictable stereochemistry without invoking carbocation rearrangements.
Introduction
Peracid/acidic hydrolysis sequences convert alkenes into anti-1,2-diols in two steps. First, a peracid (RCO₃H) delivers an electrophilic oxygen to the alkene, building an epoxide and the corresponding carboxylic acid. Protonation by H₃O⁺ activates the epoxide toward backside attack. Water, acting as the nucleophile, opens the protonated epoxide by backside attack; a tertiary epoxide carbon is generally favored over a secondary one. Deprotonation by water completes the anti diol. The illustrated hydrolysis does not require a free carbocation or rearrangement. Whether a meso product or an enantiomeric pair forms depends on the substrate geometry and symmetry.
Quick Summary
- Reagents: Peracid (e.g., mCPBA or RCO₃H) followed by aqueous acid (H₃O⁺).
- Outcome: Anti vicinal diol; an OH group occupies each original alkene carbon.
- Mechanism: Concerted epoxidation → protonated oxonium → anti water attack → deprotonation.
- Stereochemistry: Epoxidation is stereospecific (cis → cis, trans → trans); acidic opening inverts the attacked center. Cycloalkenes give trans-1,2-diols; acyclic substrates can give meso products or enantiomeric pairs.
- Rearrangements: A free carbocation and shifts are not required in the standard aqueous hydrolysis mechanism.
- Common pitfalls: Forgetting to protonate the epoxide, drawing syn addition, or omitting the final deprotonation.
Mechanism (Anti Dihydroxylation)
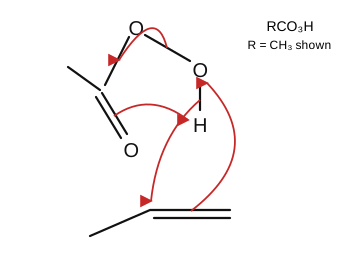
Step 1 — Epoxidation. The alkene undergoes Prilezhaev epoxidation: a concerted “butterfly” set of curved arrows delivers the terminal peroxy oxygen (Ot) to the C=C, breaks the O–O σ bond, transfers the acidic proton to the carbonyl oxygen, and closes the three-membered epoxide ring as the carboxylic acid forms. The peracid leaves as the corresponding carboxylic acid, and alkene geometry is retained (cis → cis, trans → trans).

Step 2 — Protonation. Hydronium donates a proton to the epoxide oxygen, generating an oxonium ion. Protonation increases the electrophilicity of the ring carbons and allows C–O bond cleavage to leave a neutral OH group on the other carbon.

Step 3 — Nucleophilic attack. In the representative panel, water approaches from the backside (anti) relative to the epoxide oxygen and attacks the more substituted carbon. Acid-opening regioselectivity is substrate-dependent; it does not change which carbons bear OH in the final unlabeled diol. The ring opens in an SN2-like fashion, guaranteeing inversion at the attacked carbon and establishing the anti relationship between the two oxygens.

Step 4 — Deprotonation. A second water molecule deprotonates the oxonium, yielding the neutral vicinal diol. Ring opening occurred by anti attack; an acyclic product can subsequently rotate about its C–C bond.

Step 5 — Product view. The representative propene sequence gives racemic propane-1,2-diol (connectivity shown). For substrates with two new stereocenters, determine meso versus racemic products from the alkene geometry and product symmetry.
Mechanistic Checklist (Exam Focus)
- Draw the peracid transfer as a cyclic, concerted step that forms the epoxide and carboxylic acid simultaneously.
- Protonate the epoxide before water attacks; the oxonium is the electrophile.
- Show backside attack by water (SN2-like, anti opening); a tertiary epoxide site is generally favored over a secondary site.
- Include the deprotonation that delivers the neutral diol.
- Highlight anti addition and put OH on both original alkene carbons.
- Do not introduce carbocation shifts into the standard backside hydrolysis mechanism.
Worked Examples
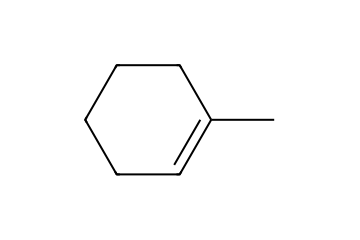
Example A — Anti Dihydroxylation of 1-Methylcyclohexene
- Substrate: 1-Methylcyclohexene.
- Reagents: RCO₃H (e.g., mCPBA), followed by aqueous acid (H₃O⁺).
- Pathway: Epoxidation → oxonium formation → anti water attack at the tertiary carbon → deprotonation.
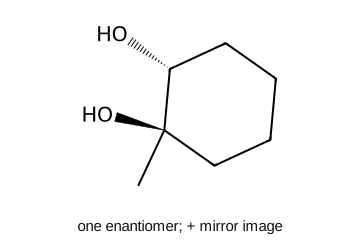
- Outcome: Racemic trans-1-methylcyclohexane-1,2-diol; incoming water supplies OH at the tertiary carbon, and the original epoxide oxygen remains on the adjacent secondary carbon. The two ring hydroxyls are trans.

When Multiple Alkenes Are Present
Peracids generally favor electron-rich alkenes, but steric accessibility and reagent amount also affect selectivity. If several sites epoxidize, hydrolysis can give several diol-containing products; substitution alone does not establish which epoxide opens fastest. Compare this pathway with alternatives such as osmylation (syn dihydroxylation) or halohydrin formation depending on the substrate and stereochemical goals. For related anti additions that install different heteroatoms, see the bromohydrin formation guide.
Practical Tips & Pitfalls
- Two-step control: Isolate the epoxide or run the sequence one-pot; ensure an acidic aqueous workup to drive ring opening.
- Solvent: Conduct epoxidation in an inert solvent (e.g., CH₂Cl₂), then add dilute acid (H₂O/H₃O⁺) for the opening step.
- Temperature: Maintain 0–25 °C to avoid side reactions and to preserve stereochemistry.
- Alternative nucleophiles: In the presence of alcohols or halides, the protonated epoxide can be opened by those nucleophiles instead of water — adjust conditions accordingly.
- Reagent handling: mCPBA and other peracids are oxidizing; store cold and neutralize residues carefully.
- Quench: After epoxidation, aqueous base (NaHCO₃) can remove benzoic acids before the acidic hydrolysis stage.
- Regiochemistry reminder: Basic opening favors the less hindered epoxide carbon. Acidic opening depends on substitution and the nucleophile, with tertiary sites often favored. Both are backside openings; hydroxide/water still gives OH at both original alkene carbons.
Exam-Style Summary
- Draw the peracid epoxidation, protonated epoxide, and anti SN2 opening.
- Anti addition gives trans ring diols; acyclic diols can be meso or racemic depending on alkene geometry.
- Track oxygen origin separately from product connectivity: the final diol has OH on both former alkene carbons.
- For the standard aqueous hydrolysis, draw backside ring opening rather than hydride or alkyl shifts.
- Compare with syn dihydroxylation (OsO₄) and bromohydrin formation to emphasize stereochemical contrasts.
Interactive Toolbox
- Open the Reaction Solver for additional practice. This general link does not preload the example; automatic handling of this sequence has not been verified.
- Drill regiochemistry choices vs. hydroboration and oxymercuration using spaced-repetition decks.
- Use mechanism builders to visualize epoxide opening pathways with different nucleophiles.
FAQ / Exam Notes
- Does the order matter? Yes. Perform the epoxidation before introducing strong acid; otherwise the alkene may be protonated instead of epoxidized.
- What if base is present? Basic opening is also an anti, SN2 backside process and generally attacks the less hindered carbon. It does not give syn addition.
- Can other nucleophiles open the epoxide? Alcohols, halides, or carboxylates can substitute for water, yielding ether or halo-alcohol products; control the solvent to favor diols.
- Are meso products possible? Yes. Anti dihydroxylation of trans-2-butene gives meso-butane-2,3-diol, whereas cis-2-butene gives the (2R,3R)/(2S,3S) pair. Oxygen origin does not distinguish two ordinary OH groups.